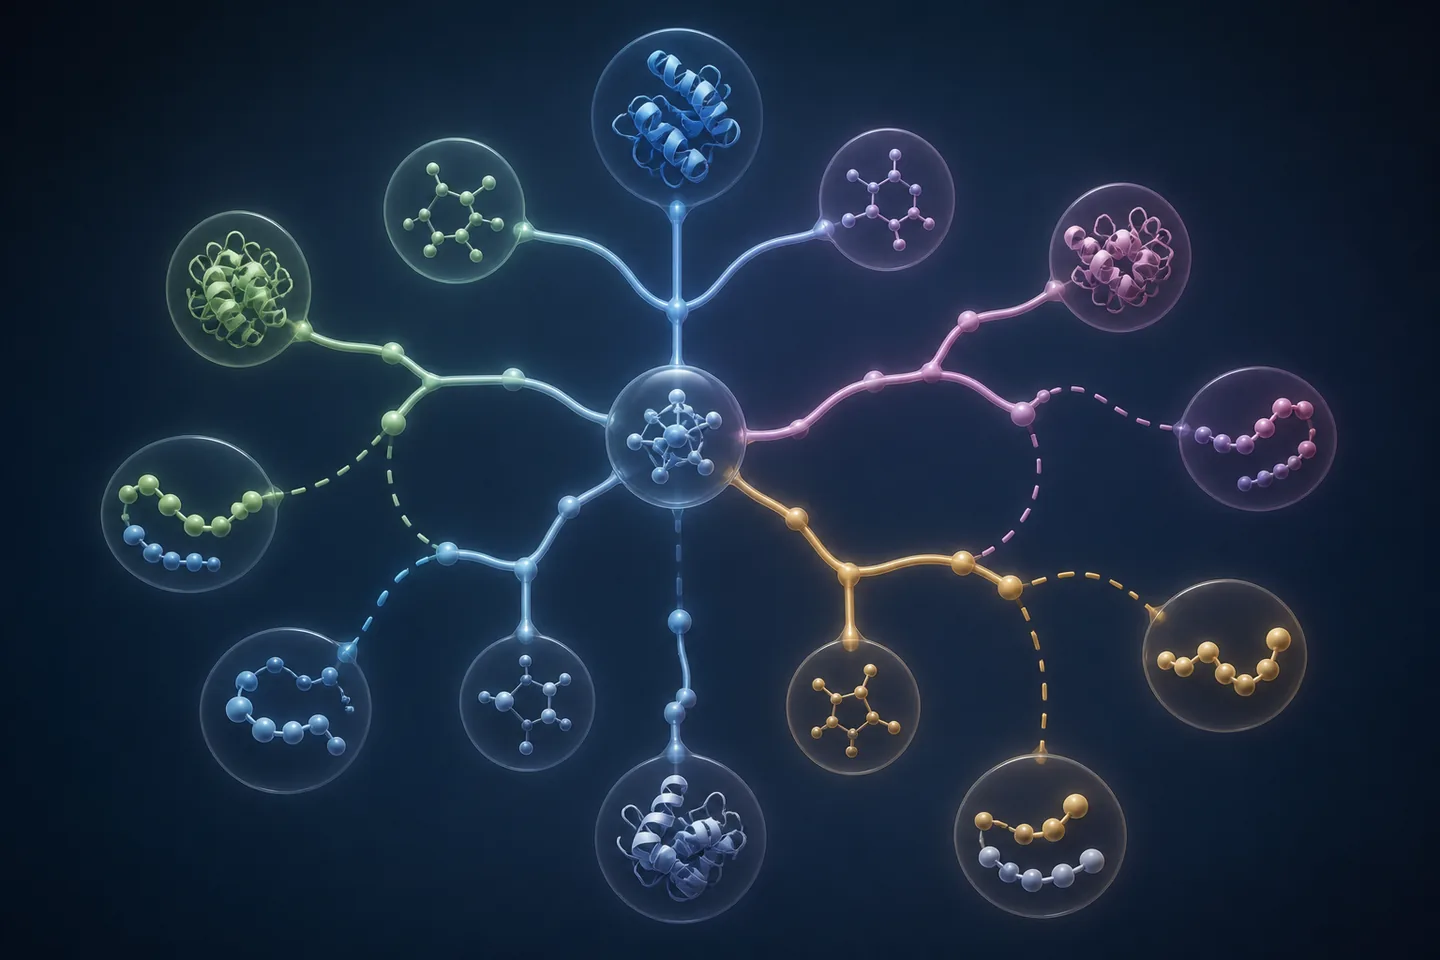
Researchers at Japanese universities and institutes have introduced ChemTSv3, a framework for generative molecular design. Conventional methods usually choose one representation in advance, thereby fixing the space they can explore. ChemTSv3 instead treats representations as nodes and molecular edits as transitions between them.
This common formulation can combine chemical strings, molecular graphs and protein sequences with different editing operations. In computational tasks, the authors show that the strategy can switch during a search and span designs from drug-like small molecules to proteins.
This is a peer-reviewed methods and computational study. It does not mean the system discovered a clinically validated medicine, and it does not by itself establish synthesis, toxicity or biological activity. Several authors are affiliated with MolNavi; the paper states that the authors declare no competing interests.
The approach could support more adaptive workflows in which chemists change objectives and representations as evidence accumulates. Independent comparisons, laboratory testing of candidates, reproducibility and computational-cost measurements are still needed.
If performance carries beyond the reported benchmarks, broader research pilots could appear within 1–3 years. Any effect on real drug or materials development would be judged over roughly 5–10 years and cannot be promised by this study.
Be the first to open the discussion.